Predicting binding affinity doesn’t work — or does it?

webinar

Tue, 16 Jun 2015, 19:00 CEST (Berlin)

Dr. Carsten Detering, BioSolveIT GmbH, St. Augustin, Germany

Predicting binding affinity doesn’t work — or does it?

Predicting binding affinity is still regarded as the holy grail. With SeeSAR, however, we are one step closer to accurately and reliably predict a protein-ligand’s binding affinity. What one often neglects is the accuracy of the data, and that, especially in brute force correlation analysis, we might compare apples with oranges.

SeeSAR will tell you where the problems are. So you can understand, without looking at any numbers, where you might have to optimize or where a crystal structure might not be credible. Because they are not, as Derek Lowe once put it, a “message from God”, but full of assumptions themselves. If we start to understand incorporate this in our work, we will see calculated binding affinity in a whole new (green) light.
Current news

VAST Just Got Bigger — The September 2026 Update
September 17, 2026 09:00 CEST
More compounds. More chemistry. More opportunities to discover your next active. This August, explore Enamine’s largest compound catalog with infiniSee xREAL as BioSolveIT’s Tool of the Month. XtalPi has expanded its VAST Chemical Space, giving you access to almost six billion virtual product molecules built from over 25,000 carefully selected...
Read on
category
Events
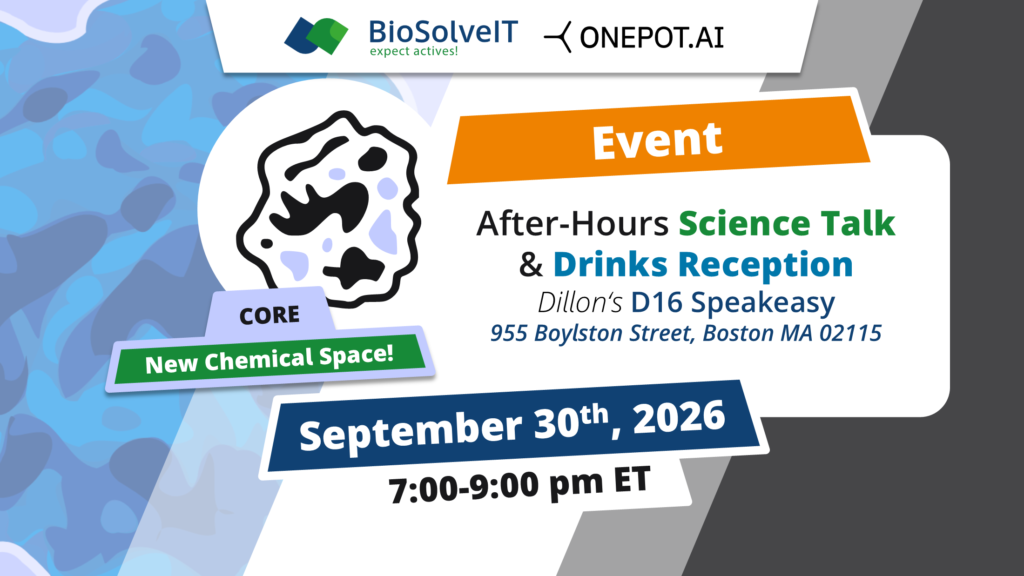
Boston Event: Expanding Molecular Diversity with CORE
September 15, 2026 17:27 CEST
Chemical Space Exploration Meets AI in Boston BioSolveIT and Onepot.AI invite you to an evening of science, drinks, and conversation at D16, the speakeasy beneath Dillon’s in Boston. Join us on September 30, 2026, for two very concise presentations exploring how ultra-large Chemical Spaces and artificial intelligence can open new...
Read on
category
Events
Beyond the Flatland: sp³-Rich Fragments Unlock Novel Kinase Chemotypes: Free Webinar!
September 15, 2026 10:40 CEST
Kinases are among the most intensively pursued target families in drug discovery, yet the chemical matter screened against them remains surprisingly narrow. Conventional kinase-focused libraries are dominated by flat, heteroaromatic compounds, while saturated, three-dimensional molecules stay largely underexplored. That narrow structural diversity makes it harder to find genuinely novel chemotypes,...
Read on