



More chemistry. Trillions of possibilities.
AMBrosia enters its third version with more than 2.4 trillion virtual product molecules. Developed in close collaboration with Ambinter (a brand of Greenpharma), the expanded Chemical Space combines 313,777 building blocks through 54 chemical reactions, implemented as 131 technical reactions, opening up a wealth of chemistry for your drug discovery projects.
AMBrosia October Update at a Glance
- 50+ chemical reactions
- 300,000 building blocks
- 2,440,635,639,301 virtual product molecules
Changes in this Update
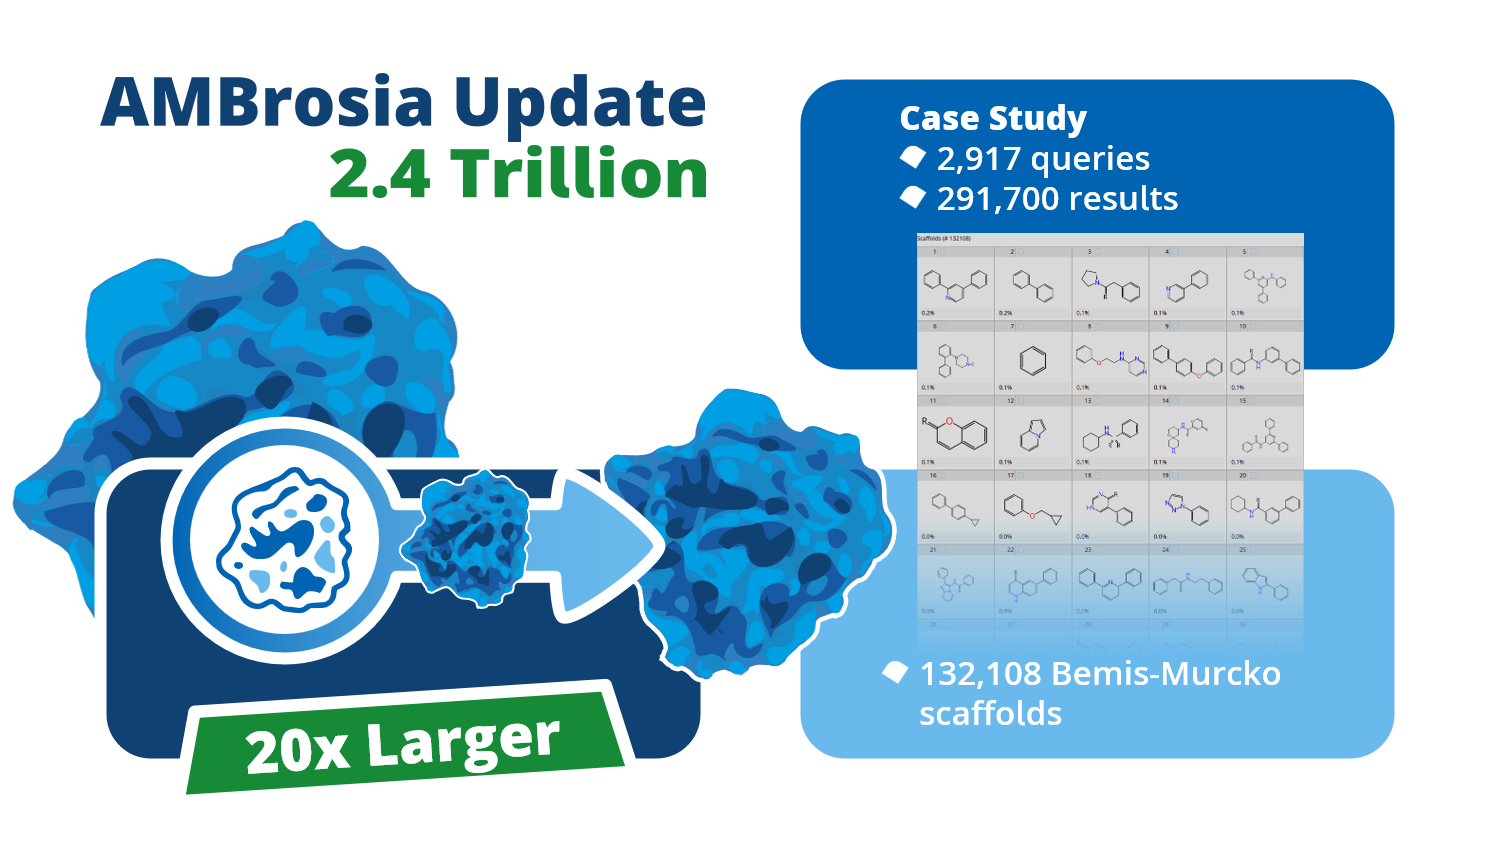
- ~20-fold increase in virtual products
- ~5× more building blocks
- 12 new reactions
Growing in Size and Chemical Diversity
From 110 billion virtual compounds at its launch in February 2024 to 126 billion in June 2025, AMBrosia now reaches more than 2.4 trillion entries. Together with Ambinter, we are extremely proud of this development. The expanded selection of building blocks and reactions enriches the Space’s chemical diversity, giving researchers more opportunities to discover relevant analogs and explore alternative scaffolds.
Case Study: Investigating Scaffold Diversity in AMBrosia
As part of AMBrosia’s quality assessment, we searched a benchmark set of 2,917 bioactive compounds (Neumann et al.) using SpaceLight with ECFP4 fingerprints, retrieving 100 results per query. Across the 291,700 retrieved results, we identified 132,108 distinct Bemis–Murcko scaffolds, highlighting the breadth of scaffold diversity in the result set.
For 706 queries, more than 50 of the 100 results achieved an ECFP4 Tanimoto similarity above 0.5. These findings demonstrate substantial coverage of similar chemistry for these queries, alongside broad scaffold diversity across the retrieved compounds.
How to Explore AMBrosia
Explore AMBrosia with infiniSee through three complementary search modes: Scaffold Hopper for pharmacophore-guided similarity, Analog Hunter for fingerprint-based analog searches, and Motif Matcher for substructure-focused exploration. The corresponding command-line tools (FTrees, SpaceLight, and SpaceMACS) also support integration into your own workflows.
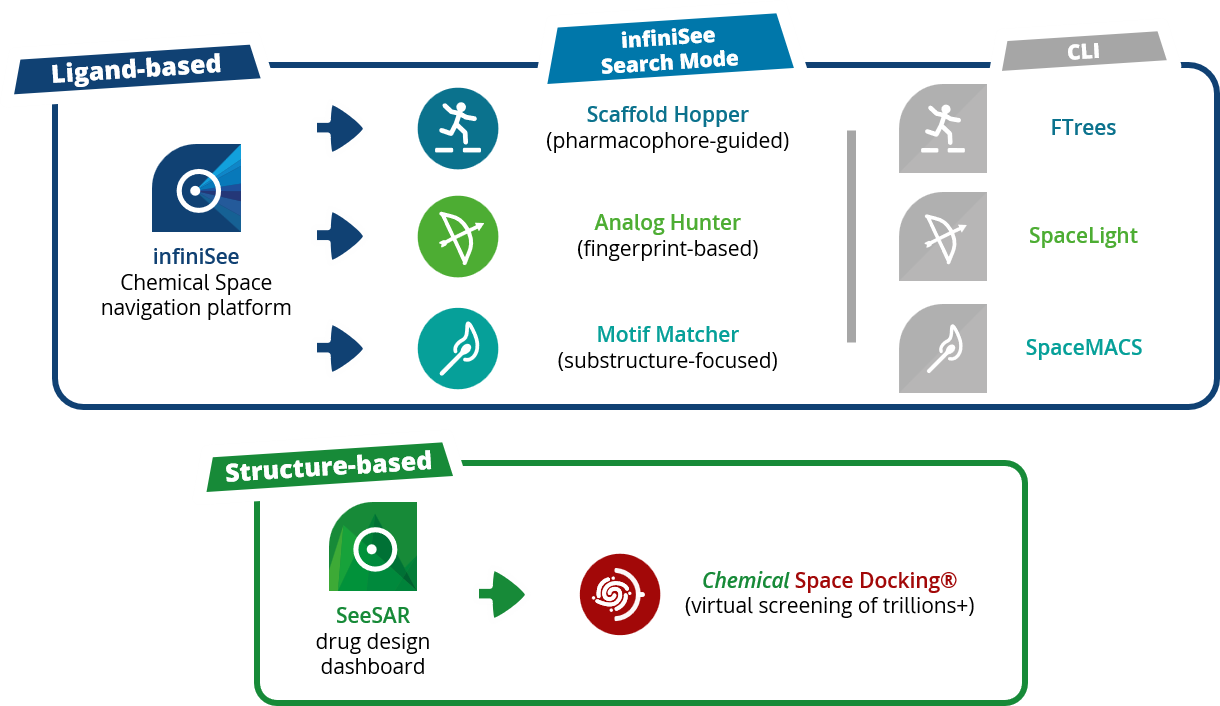
For structure-based discovery, screen AMBrosia directly within your target’s binding site using Chemical Space Docking® in SeeSAR to identify promising candidates with favorable binding-site interactions.
From Virtual Exploration to Tangible Chemistry
Building blocks are selected using BioSolveIT filter rules. AMBrosia’s reaction-based architecture links retrieved products to their constituent building blocks and transformations, providing a chemical foundation for discussions with Ambinter about synthesis. To request a synthesis quotation, contact ambrosia@greenpharma.com with your selected structures, compound IDs, and required quantities.
Discover the expanded AMBrosia Space and expect actives!